Tautomer Search
Python API · stjames models · API example
How it works
Predict the relative free energies and populations of a molecule's tautomers. Rowan enumerates candidates, screens out high-energy forms, and refines the conformer ensembles of the remaining tautomers.
Settings
- Conformer sampling: "Conformer generator" (
conf_gen_settings) defaults to openconf, with "Max conformers" set to 20 per tautomer. ETKDG, iMTD (CREST), and MCMM are also available. Openconf defaults to 200 steps and a 10 kcal/mol conformer energy window; advanced controls are covered in Conformer search. - Energy method: "Optimization method" (
multistage_opt_settings) controls geometry refinement and final single-point scoring. The web default optimizes with AIMNet2/ALPB(water) and scores with AIMNet2/CPCM-X(water). The g-xTB preset optimizes with gas-phase GFN2-xTB and scores with g-xTB/CPCM-X(water). "Custom" lets you edit methods and solvents for individual stages; "Solvent (all steps)" applies a common solvent to supported stages. See Multistage optimization. - Tautomer screening: "Screening window (kcal/mol)" (
screening_window, default 10) controls the maximum relative energy retained during the initial screen. A wider window carries more candidates into refinement; at most 50 tautomers proceed. This is separate from the conformer energy window. - Solvation correction: "Final correction" (
final_correction) defaults to "None." "COSMO-RS" ("COSMO_RS"in the API) adds a more expensive solvation free-energy correction. Its solvent follows the final single-point solvent, defaulting to water if none is specified.
For reproducible API submissions, pass the conformer and optimization settings explicitly. Omitted settings use the installed stjames defaults, which can differ from the web presets.
Notes
Enumeration includes same-net-charge protonation microstates, including zwitterions that RDKit's tautomer rules can miss. The search keeps the input net charge fixed, so it does not predict a pH-dependent distribution across charge states.
The initial screen uses GFN2-xTB geometry optimization and the selected scoring method. Refinement then combines conformer energies with vibrational thermal corrections into one ensemble free energy per tautomer. With COSMO-RS, only the three lowest-free-energy conformers before that correction contribute; the selected scoring method's electronic energy remains the basis for ranking.
Results include ranked structures, relative free energies in kcal/mol, and Boltzmann populations at 298.15 K. Populations describe the successfully calculated candidates under the chosen settings. Close rankings can be sensitive to sampling and solvent treatment; inspect competing forms before choosing one for downstream work.
Submission video
Benchmarks and validation
Accuracy
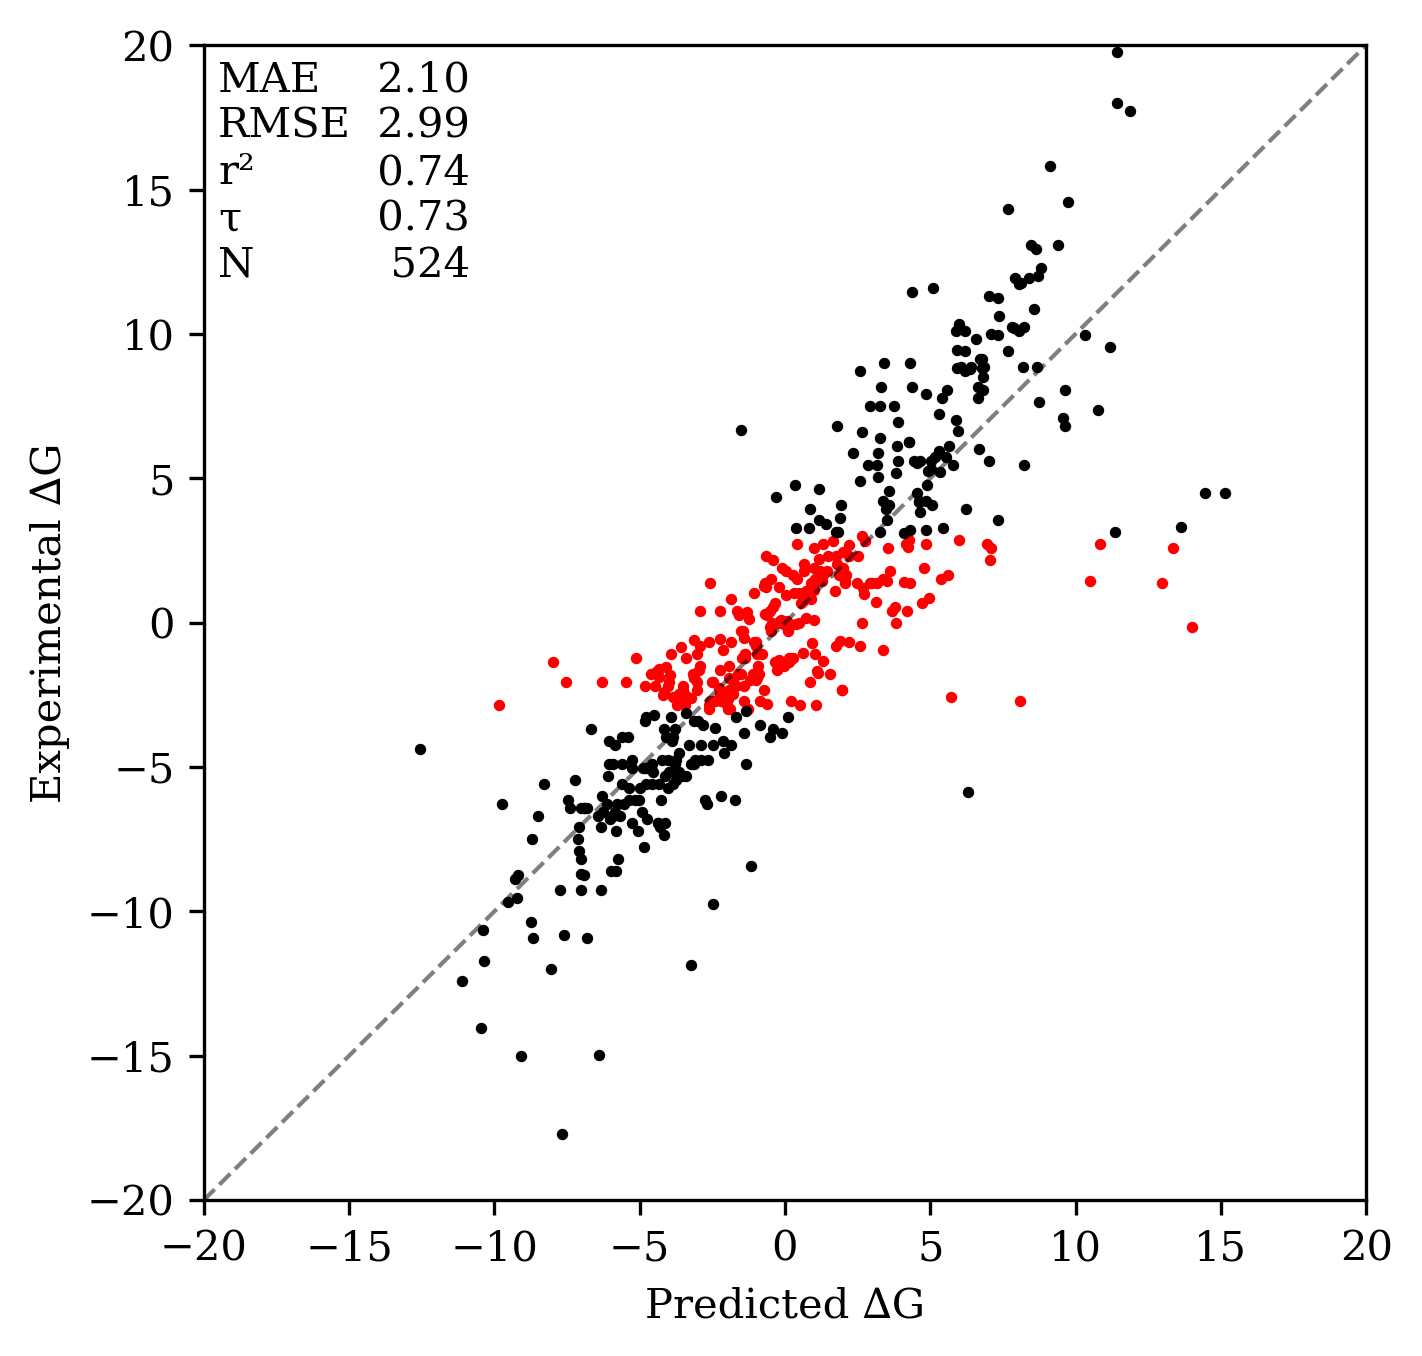
This historical benchmark used the old default protocol: ETKDG conformer sampling and AIMNet2/CPCM-X(water) single-point energies on GFN2-xTB geometries. These results do not validate the current default settings or the optional COSMO-RS correction.
On the aqueous subset of the TautoBase benchmark set, Rowan's tautomer workflow displays a mean absolute error of 2.10 kcal/mol and a root mean squared error of 2.99 kcal/mol. This is comparable to the performance of high-level quantum chemical methods reported by Chodera and co-workers: B3LYP/aug-cc-pVTZ with B3LYP/6-31G(d)/SMD solvation gave an RMSE of 3.1 kcal/mol on a different TautoBase subset.
The workflow identified the correct lowest-energy tautomer in 89% of the benchmark cases. For the closer comparisons, with an experimental free-energy difference below 3 kcal/mol (shown in red), classification accuracy was 77%.
Rowan's tautomer search workflow's performance on TautoBase